Chapter 5
Many-Electron Atoms
5.1Introduction
When a second electron is added to hydrogen, the problem ceases to be solvable in closed form. This is not merely a matter of difficulty: the situation is the same as that of the three-body problem in classical mechanics, and no exact expression exists for the ground-state energy of helium.
The character of the subject therefore changes at this point. From here on we work with controlled approximations, and the essential skill is to judge how good each one is. Helium is a convenient case in which to develop that skill. Three successively better treatments, each requiring about a page of algebra, bring us to within two percent of the true answer, and each of them introduces an idea that is used repeatedly later.
Four results are established in this chapter:
Antisymmetry, from which the Pauli principle follows as a theorem rather than a postulate;
Screening, which we compute rather than assume, and which breaks hydrogen's accidental -degeneracy and produces the quantum defect;
Exchange, an energy splitting of nearly an electron volt that arises from antisymmetry alone and has no classical analogue;
Term symbols, the notation in which tables of atomic data are written.
The chapter closes with a state that can decay without absorbing radiation, namely a bound state degenerate with a continuum. Such states are the subject of the final chapters of this book, and helium provides the simplest example of one.
Read antisymmetry (Sec. 5.2), the helium Hamiltonian (Sec. 5.3), and then the three successive treatments of the same atom in Secs. 5.4, 5.5 and 5.6, which are the core of the chapter: one problem solved three times, each time better, with the reason for the improvement visible. Add exchange (Sec. 5.7), then work the three self-checks in Sec. 5.10.1 before the Summary. Screening and the quantum defect (Sec. 5.8) are needed for Chapters 8–11 and for the companion course Quantum Chemistry; term symbols (Sec. 5.9) are notation to consult rather than to learn in one pass; autoionization (Sec. 5.10) is the seed of Chapters 10 and 11 and may be postponed.
Chapter 1 minimized the Rayleigh quotient over the two components of a vector, and the minimum was the exact lower eigenvalue because the two-dimensional trial space was the whole space. This chapter minimizes the same quotient over a one-parameter family of functions inside an infinite-dimensional space, so the minimum is an upper bound rather than the answer. Nothing about the principle changes; only the size of the search space does, and with it the meaning of the number that comes out. That distinction — exact diagonalization in a small space versus optimization within a restricted family — is the whole of what separates the methods catalogued in Quantum Chemistry.
5.2Identical Particles and Antisymmetry
Electrons are indistinguishable, in the sense that no measurement can determine which is which. Formally, the exchange operator that swaps particles and must commute with the Hamiltonian, and since its eigenvalues are . Fermions, and electrons among them, take the eigenvalue .
The state of a system of identical fermions changes sign under exchange of any two of them:
where collects position and spin.
A postulate, not an axiom, and the difference matters here. The six axioms of Chapter 1 say what quantum mechanics is; nothing in them forces the choice of sign above. Both eigenvalues of are allowed by the formalism, and the symmetric choice describes a perfectly consistent world — one made of bosons. We take the antisymmetric one for electrons because that is what the world does, and the periodic table is the evidence.
No two identical fermions can occupy the same one-particle spin-orbital.
Setting in Eq. (5.1) makes the left- and right-hand sides the same function, so that and therefore . A state with two electrons in the same spin-orbital does not exist.
The exclusion principle is thus not an additional rule imposed on quantum mechanics; it is a two-line consequence of antisymmetry.
For electrons occupying spin-orbitals , the normalized antisymmetric state is
This construction works because a determinant changes sign when two rows are interchanged, which is Eq. (5.1), and vanishes when two columns are equal, which is Theorem 5.2.1.
Write out Eq. (5.2) for and verify both properties.
Solution.
For two electrons,
Antisymmetry. Swapping gives
Exclusion. Setting ,
Normalization. If and are orthonormal, then expanding gives four terms: two of the form , each with prefactor , and two cross terms proportional to . Hence , which is the reason for the prefactor .
5.3Helium: The Hamiltonian
Helium consists of two electrons and a nucleus of charge with . Before writing a single symbol, it is worth drawing the object we are about to describe, because the drawing already contains the whole difficulty of the chapter. There are exactly three distances in this system. Two of them run from the nucleus to an electron. The third runs between the two electrons, and it is the one that will resist every method in this chapter.
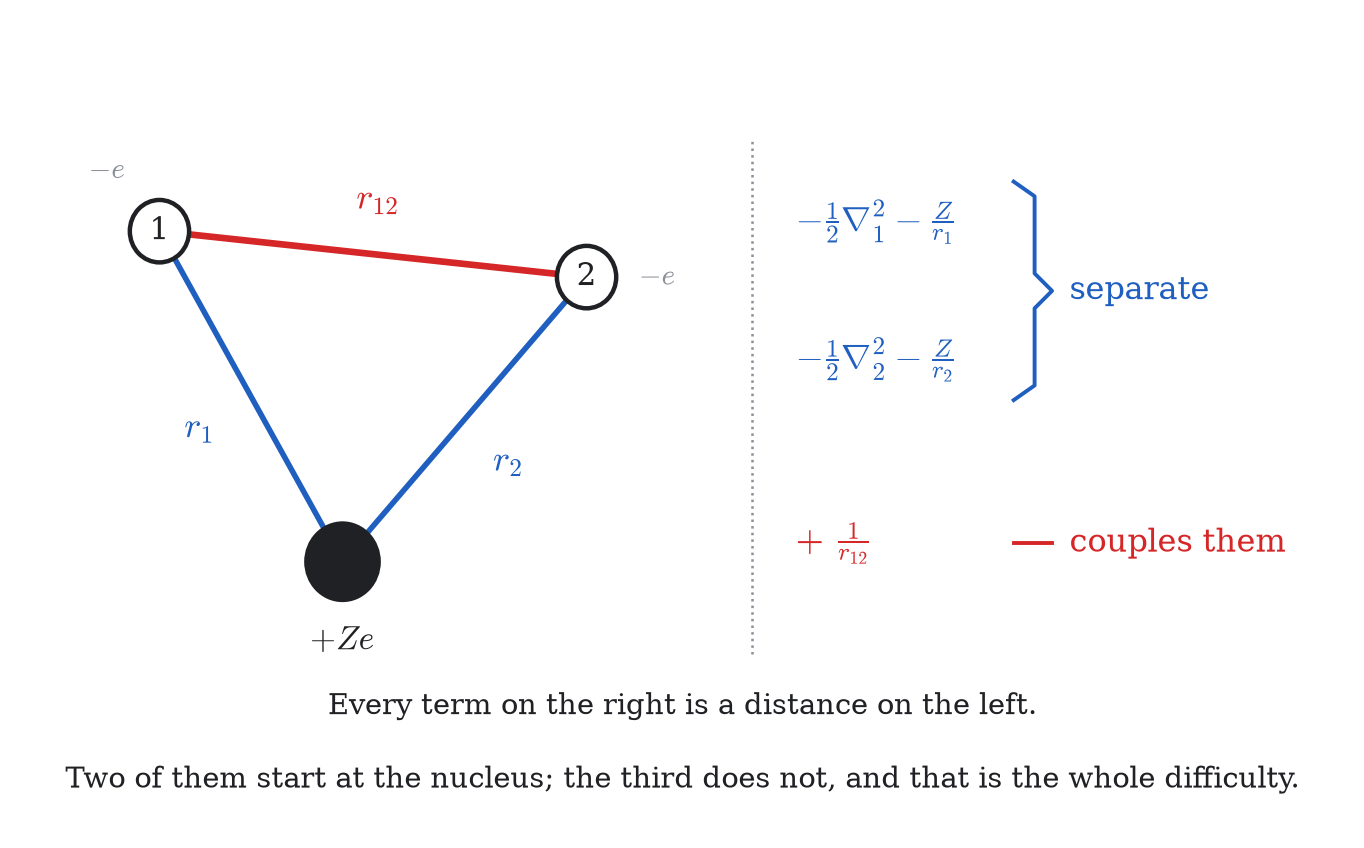
Working in atomic units (Chapter 4, Sec. 4.8), the Hamiltonian is
with . In SI units the last term is .
The operators and each involve one electron only, so that if were absent the equation would separate and the solution would be a product of two hydrogenic orbitals with . The term couples the coordinates. It cannot be written as , no change of variables removes it, and consequently no separable solution exists. This single term is the source of essentially all the difficulty in many-electron atoms.
The experimental values to be reproduced are
5.4First Approximation: Independent Electrons
We begin by dropping entirely. The equation then separates, and each electron occupies a hydrogenic orbital with ,
which is from Chapter 4, Eqs. (4.52) and (4.32), collected into a single expression.
Compute the ground-state energy of helium neglecting the electron–electron repulsion.
Solution.
Each electron has the hydrogenic energy of Eq. (4.48) with ,
so that for two independent electrons
Compared with the true value of eV this is too low by eV, an error of . The sign of the error is instructive. We have omitted a repulsion, and a repulsion raises the energy, so neglecting it must give a value that is too negative. An approximation whose error has the wrong sign therefore contains a mistake.
5.5Second Approximation: First-Order Perturbation Theory
We now treat as a perturbation and add its expectation value in the zeroth-order state:
This is a six-dimensional integral, but it can be evaluated exactly, and the method is worth following in detail because the same device is used again later.
For the normalized hydrogenic orbital of Eq. (5.5),
The charge density of electron , , is spherically symmetric. By the classical shell theorem, the potential it produces at radius separates into a contribution from the charge inside, which acts as if concentrated at the origin, and one from the shells outside, each of which contributes :
Inner integral. With and the standard result with ,
Outer integral. With ,
Combining. Dividing the first by and adding,
The terms cancel, which provides a useful check.
Averaging over electron 1. We then have
This splits into three elementary integrals, evaluated with :
Therefore
Use Theorem 5.5.1 to correct Example 5.4.1.
Solution.
For ,
so that
The estimate has moved from too low to too high. This overshoot is to be expected, since the orbitals were not allowed to respond to the repulsion. Each electron is therefore held closer to the nucleus than it actually is, and experiences more repulsion than it actually does. Allowing the orbitals to relax is the subject of the next approximation.
5.6Third Approximation: The Variational Method
For any normalized trial state and any Hamiltonian with ground-state energy ,
Expand in the exact eigenstates, with and . Then
since every .
The principle converts approximation into optimization: we choose a trial state containing adjustable parameters, compute , and minimize the result. The answer obtained is guaranteed to be an upper bound, which is a useful property in practice, since a calculation that returns an energy below the exact value must contain an error.
The physical shortcoming of Sec. 5.5 was that each electron still experienced the full nuclear charge , although its partner partly shields it. We therefore allow the orbital exponent to be a free parameter :
Minimize over and evaluate the result for .
Solution.
Three expectation values are needed. For a hydrogenic orbital with exponent , Chapter 4's results Eq. (4.59) give , and the virial relation for that orbital gives .
Kinetic energy, two electrons:
Nuclear attraction, two electrons, with the true charge in the operator but the trial exponent in the state:
Electron repulsion. Theorem 5.5.1 was derived for orbitals of exponent ; here the exponent is , so that
Adding these,
Minimizing,
Substituting back, and noting that ,
For helium, :
The error is now , and, as Theorem 5.6.1 requires, the result lies above the exact value of eV.
The intermediate result carries the physics of the calculation. An electron in helium does not experience the full charge ; it experiences , its partner having screened away about of an elementary charge. This idea, that the nuclear charge an electron actually experiences is smaller than , organizes the remainder of the chapter.
The three treatments give, in order, , and eV, against the true value of eV. This pattern is characteristic. A crude model can be badly wrong; first-order perturbation theory usually overcorrects; and allowing the wave function itself to relax recovers most of the remaining discrepancy. Going further requires trial functions that depend on explicitly, which is where quantum chemistry proper begins and where this course stops.
5.7Exchange: Singlet and Triplet Helium
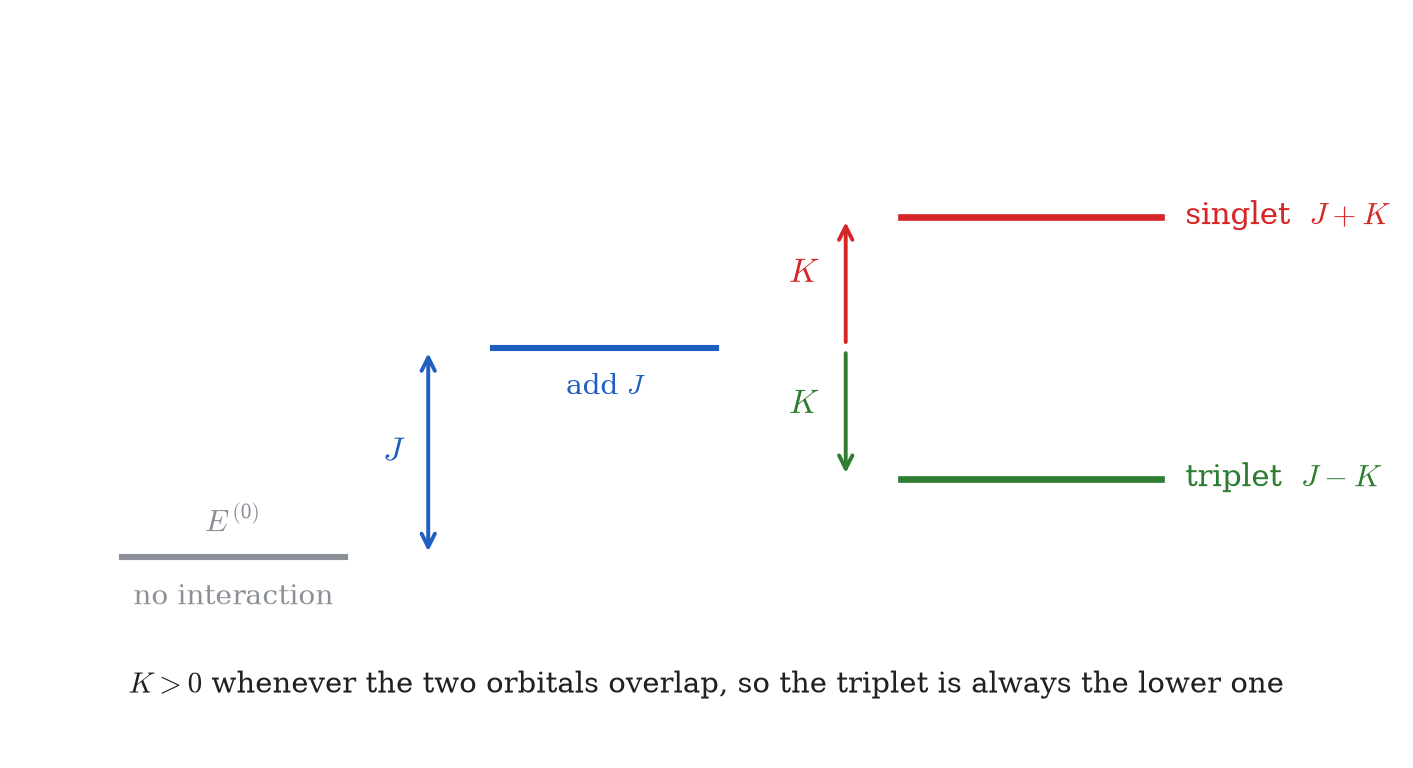
We now excite one electron, to the configuration , and denote the two spatial orbitals by and . Antisymmetry of the total state can be achieved in two ways, because the state factorizes into a spatial part and a spin part, and it is sufficient that one of the two be antisymmetric.
Both possibilities are physically admissible, and their energies differ. The calculation below shows precisely why.
The integral admits a classical interpretation: it is the electrostatic repulsion between two charge clouds and . The integral admits none, since the orbitals are interchanged between the two factors and no classical charge distribution corresponds to such an expression. It arises only because the wave function must be antisymmetrized.
For the states of Eq. (5.11),
with for the singlet and for the triplet. Since for orbitals with overlapping densities, the triplet lies lower.
The one-electron parts give regardless of the symmetrization. For the repulsion, we insert Eq. (5.11) into and expand the product of the two brackets. Four terms result. Two of them have the form , each with prefactor , and together give . The remaining two are the exchange terms, each with prefactor , and together give .
The Hamiltonian (5.3) contains no spin operator, yet the two states differ in energy by and are distinguished only by their spin arrangement. The spin does not cause the splitting; antisymmetry does. The requirement that the total state change sign under exchange ties the spin arrangement to the spatial arrangement, and it is the spatial arrangement to which the Coulomb repulsion is sensitive.
The mechanism can be seen directly in Eq. (5.11). Setting makes the antisymmetric spatial function vanish identically, so that the triplet electrons avoid one another, are further apart on average, and repel less strongly. This is the exchange interaction, and it has no classical counterpart.
The levels of helium are observed at eV () and eV () above the ground state. Extract and comment on its magnitude.
Solution.
From Eq. (5.14) the separation is :
The triplet lies lower, which confirms that .
It is instructive to compare this with the magnetic interaction between two electron spins, which at a separation of a few is of order eV, four orders of magnitude smaller. The splitting is therefore not magnetic in origin. It is electrostatic, acting through antisymmetry.
The consequence is chemical as well as spectroscopic. The rule that electrons in different orbitals favor parallel spins, known as Hund's first rule, is this same , and it is the reason that the ground state of oxygen is a triplet.
The two families were at one time thought to be different substances and were named parahelium (singlet) and orthohelium (triplet); the names have survived. We note also that cannot reach the ground state by emission of a single photon, since the transition would have to change , which the dipole operator of Chapter 3 cannot do. The state is therefore metastable, with a lifetime of about two hours. This may be compared with hydrogen's state in Example 3.6.1, which is a second instance of a forbidden route producing a long-lived state.
5.8Heavier Atoms: Screening and the Quantum Defect
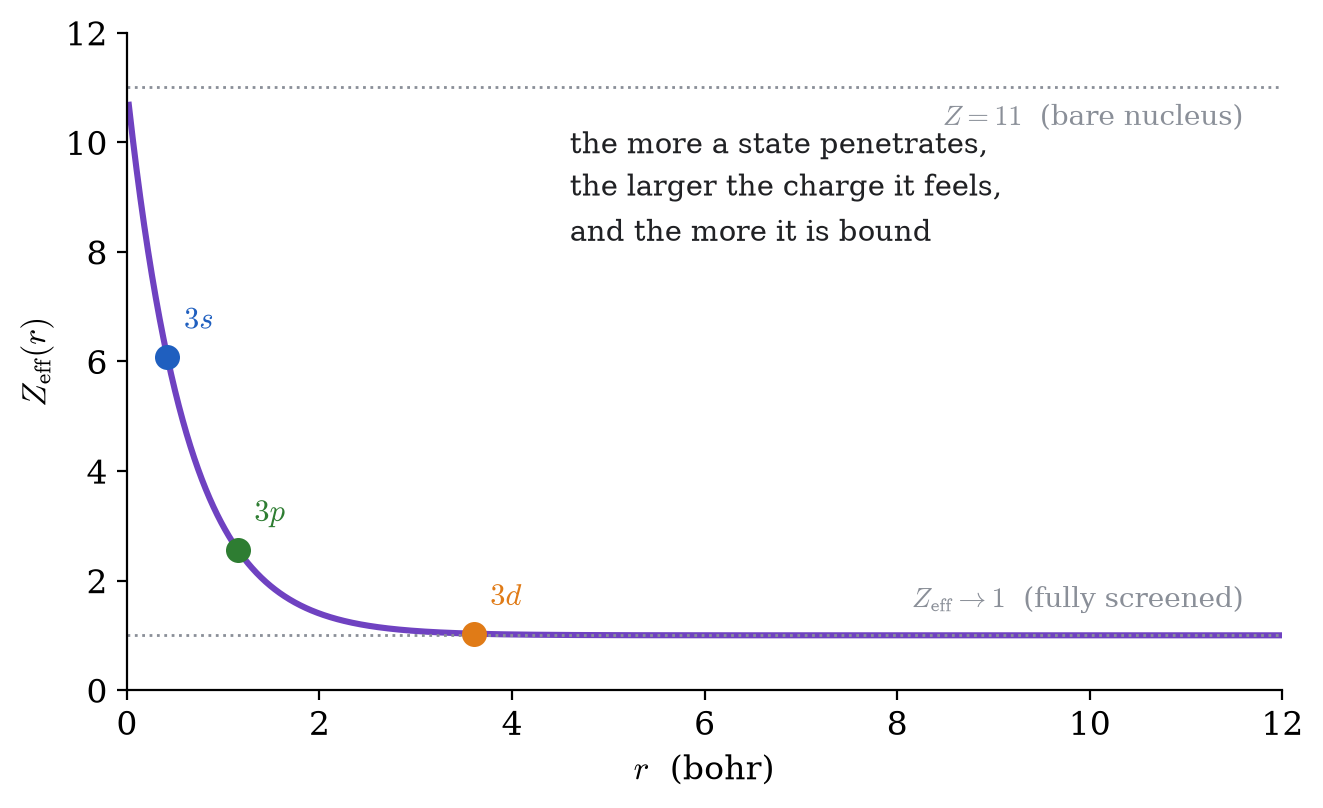
For atoms with more than two electrons, the practical approach is to retain the one-electron picture and to modify the potential. Each electron is taken to move in a central average field produced by the nucleus together with all the other electrons, so that the machinery of Chapter 3 applies unchanged: the states are labeled , and Eq. (3.44) is solved with a modified .
The essential feature of this field is its behavior at the two extremes:
An electron far from the nucleus experiences a net charge of and is hydrogenic, whereas one that penetrates the core experiences a considerably larger charge and is correspondingly more tightly bound. Chapter 3 has already established which orbitals penetrate: the centrifugal barrier in Eq. (3.44) excludes states of high from the core entirely, while has no barrier at all. It follows that
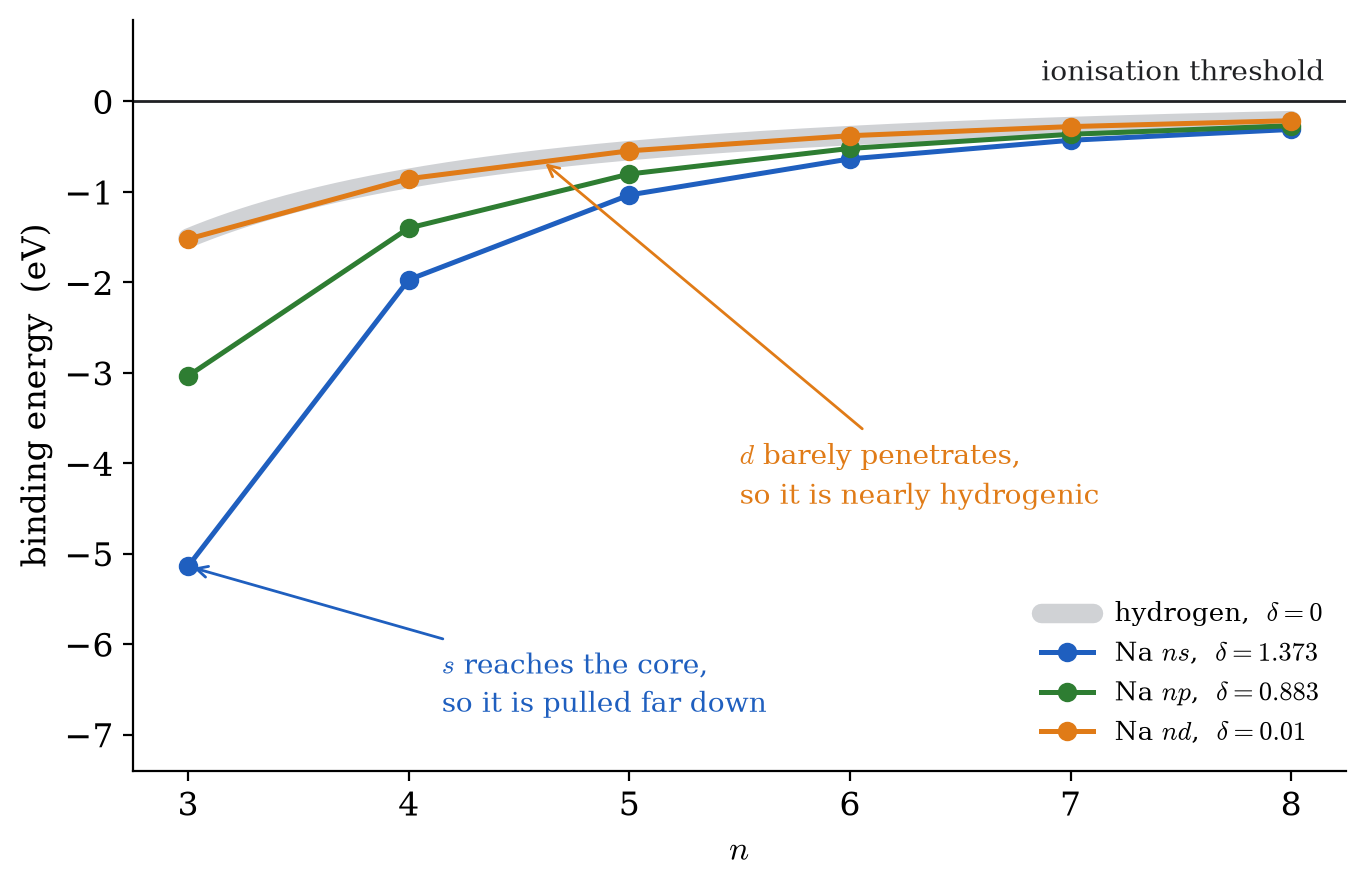
The accidental -degeneracy of Chapter 4 is therefore broken, and in a definite order. The effect is described by a single number for each value of .
The energies of a series of states of the same converging to an ionization threshold are written
with eV. The number is the quantum defect and the effective quantum number.
Two properties make this description useful. First, is nearly independent of within a series, since it is a property of the core, which is sampled only during the small fraction of the orbit spent inside it; measuring once therefore predicts every member of the series. Second, inverting Eq. (5.17) converts a measured binding energy directly into :
Sodium has one electron outside a closed-shell neon-like core, so that it is hydrogen-like in structure and serves as the standard illustration. Its ionization energy is eV above the ground state. The ground state is bound by eV, the state by eV, and the state by eV. Find and for each.
Solution.
Applying Eq. (5.18) three times,
The ordering is that of Eq. (5.16), and the magnitudes are worth examining. The electron is hydrogenic to within a third of a percent, since its centrifugal barrier holds it so far outside the core that it barely distinguishes sodium from hydrogen. The electron, with , behaves like a hydrogen state with between and , because on every orbit it passes through the neon core and briefly experiences a charge much larger than .
A single number, , together with a single mechanism, penetration, thus accounts for the structure of the alkali spectrum. The same description applies to every Rydberg series in this book, including the molecular series of Chapter 9.
5.9Term Symbols
A configuration lists which orbitals are occupied; carbon, for example, is . A configuration is not a state, since the angular momenta of the electrons can couple in several ways, giving several energies. The resulting states are labeled by a term symbol.
where is the total spin, the total orbital angular momentum written as a capital letter (), the total angular momentum obtained by coupling and , and the multiplicity ( singlet, doublet, triplet).
A filled subshell has and , and therefore contributes to a term symbol. A subshell containing one hole has the same and as one containing one electron.
In a filled subshell every from to is occupied twice, once with each spin. Hence and , and since this is the only state of the configuration, . For the second statement, removing one electron leaves and of the removed electron, so that the available values are those of a single electron with reversed signs. This is the same set, and hence gives the same and .
These two results remove most of the work. In particular, ionizing a closed-shell atom or molecule always leaves one hole and therefore always gives a doublet, .
Assign term symbols to helium's ground state, sodium's ground state, and sodium's first excited state.
Solution.
Helium, . This is a closed shell, so that by Theorem 5.9.1 and , hence and the term is . This is a singlet, consistent with Sec. 5.7, where the ground configuration admitted only the spin-antisymmetric combination.
Sodium, . All shells are closed except for one electron, so that , and : the term is .
Sodium, . Here and , so that can be or , giving the two terms and . These are split by the spin–orbit interaction, and the two transitions down to are the sodium D lines at and nm, which produce the yellow light of a sodium street lamp. The doublet structure exists only because the electron has spin.
5.10States Inside a Continuum
One further feature of many-electron atoms must be introduced, since the final chapters of this book depend on it entirely. It has no counterpart in hydrogen.
The ionization threshold of helium is eV. Above that energy the system can exist as He together with a free electron of any kinetic energy, which is to say as a continuum. Suppose now that both electrons are excited, to the configuration . That state lies at eV, well above the threshold. It is a perfectly good bound state of the two-electron system, in that both electrons are attached and it has a definite energy, and it is at the same time degenerate with the continuum.
Such a state cannot survive. The repulsion of Eq. (5.3) connects it to the continuum at the same energy: one electron drops to while the other takes up the released energy and leaves. No photon is absorbed or emitted.
Autoionization is the decay of a bound state that is degenerate with a continuum, driven by an interaction internal to the system rather than by absorption of radiation.
Because the state decays, it is not sharp: it has a width related to its lifetime by . For helium's resonance meV, so that
This configuration, a discrete state embedded in and coupled to a continuum, produces the asymmetric spectral profiles that Fano explained in 1961, using this same helium resonance as his example. We do not yet have the tools to treat it, since it requires the continuum normalization mentioned at the end of Chapter 3 and the time-dependent methods of Chapter 6. The treatment is given in full in Chapter 10.
Two points should be carried forward from this section. A doubly excited atom is a bound state inside a continuum, and it lives for femtoseconds rather than nanoseconds. The cause is the same that made the ground state of helium impossible to compute exactly. The term that could not be handled in Remark 5.3.1 thus turns out to drive the physics at which this course is aimed.
5.10.1First-Pass Self-Checks: Counting, Improving, and Missing
Two orthonormal spatial orbitals and are available to two electrons. How many antisymmetric states are there, and how do they sort by total spin?
Solution.
Two spatial orbitals carry two spin projections each, so there are four spin-orbitals. An antisymmetric state of two electrons is fixed by which two spin-orbitals are occupied — that is the content of Definition 5.2.1 — so the count is the number of ways of choosing two of four:
Sorting them by is the fastest way to identify the spins, because is fixed for each determinant while is not:
| Determinants | Count | |
| 1 | ||
| , , , | 4 | |
| 1 | ||
A triplet contributes one state to each of ; a singlet contributes one state to only. The single states at therefore belong to one triplet, which takes one of the four states with it. The remaining three states are singlets. So
and the triplet is the spatially antisymmetric combination of Eq. (5.11), lying below its singlet partner by Theorem 5.7.1.
The three singlets are the two closed-shell determinants and together with the open-shell combination, and in general the Hamiltonian mixes all three: the problem in this space is the diagonalization of one matrix, which is nothing beyond Chapter 1. Spatial symmetry can cut it down further. If and are the gerade and ungerade orbitals of a homonuclear diatomic, then and are both gerade while is ungerade, so the open-shell singlet cannot mix with the other two and the falls apart into a and a . Chapter 7 earns those labels; the point here is that the block structure comes from symmetry and not from the counting.
This is a calculation the reader will meet three more times under three different names: it is the full configuration interaction problem in a two-orbital active space in Quantum Chemistry, the Hubbard dimer at half filling in Condensed Matter Physics, and a two-qubit state space in Quantum Computing. Counting it once here is what makes those three recognizable as the same object.
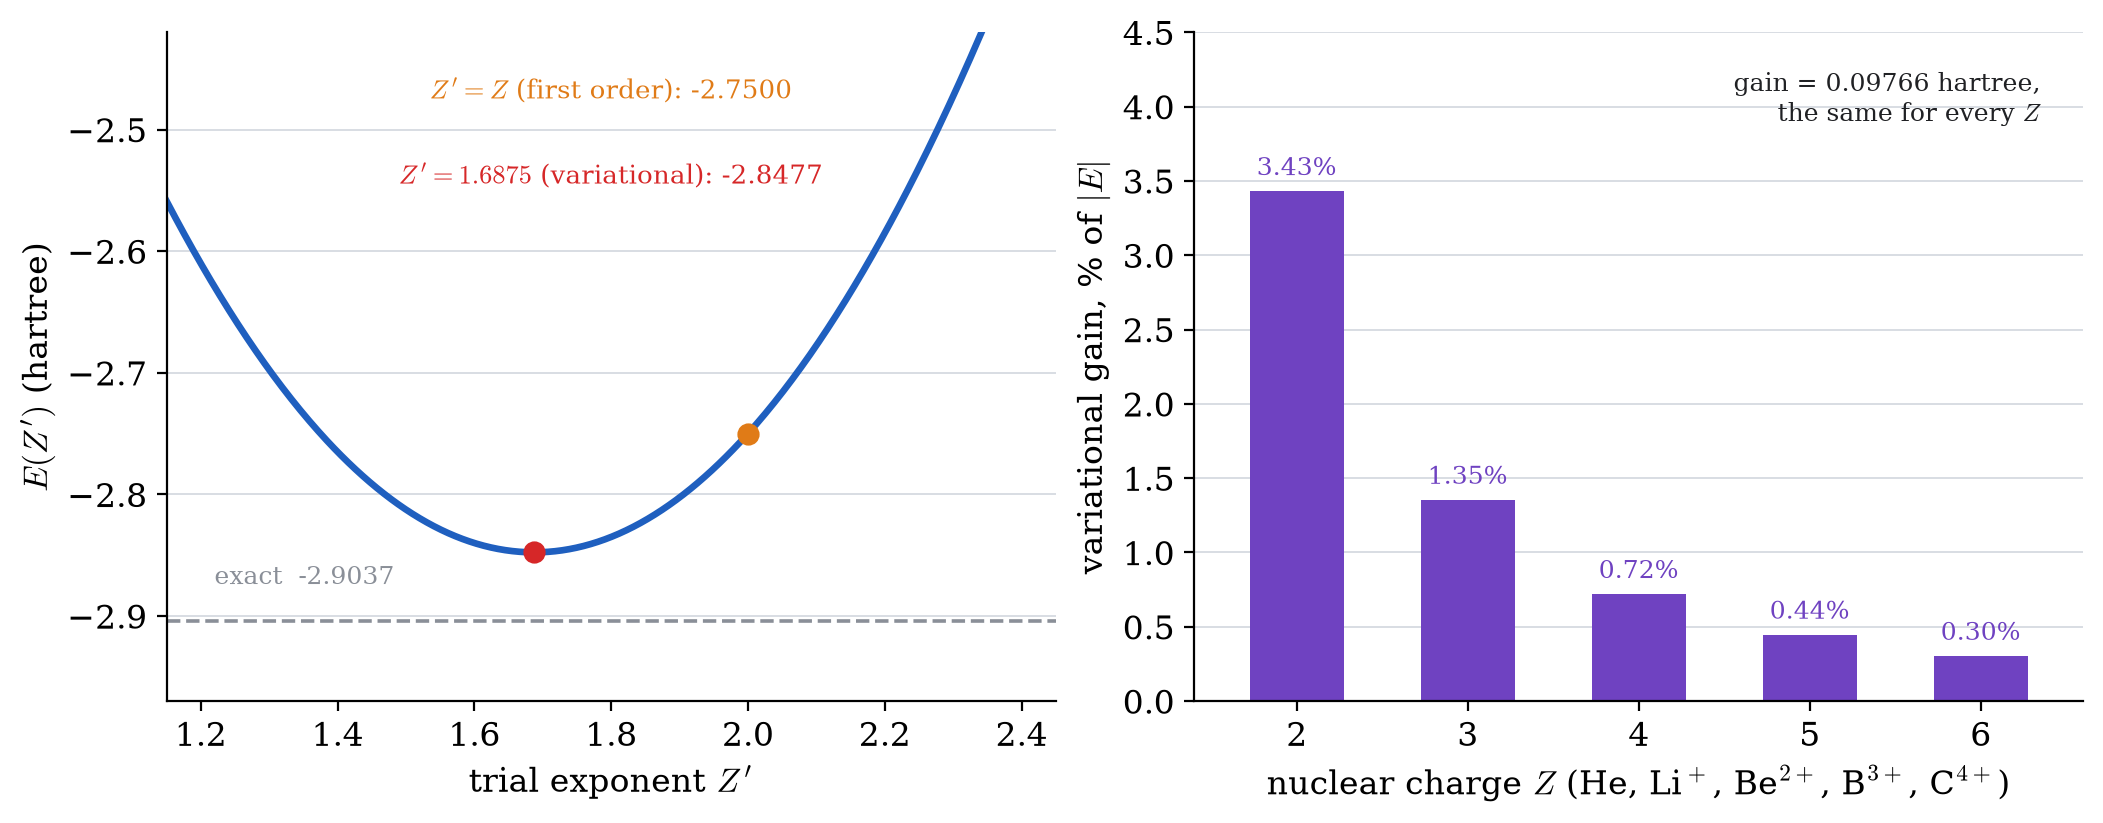
Using Eq. (5.10), compare first-order perturbation theory with the variational result for general , and evaluate the difference for He and for Li.
Solution.
First-order perturbation theory keeps the unperturbed orbitals, so it is Eq. (5.10) evaluated at :
The variational result is , which expands to
Subtracting,
independent of . Every term that depends on has cancelled.
For helium, : and a.u., so the gain is of the binding energy. For Li, : and a.u., the same a.u. but now only . Figure 5.5 carries this out to .
The physical reading is that screening is an absolute effect of fixed size, while nuclear binding grows as . Relaxing the orbital is therefore essential for neutral helium and increasingly irrelevant for highly charged ions — which is precisely the statement that the electron–electron term is a genuine perturbation only when is large.
Predict helium's first ionization energy from the variational model, and account for the difference from the true value.
Solution.
Ionizing helium leaves He, a one-electron ion, whose energy Chapter 4 gives exactly: a.u. So the model predicts
The exact non-relativistic value, quoted rather than derived here, is a.u., giving a.u. eV, in agreement with the measured eV to within the relativistic and finite-nuclear-mass corrections this chapter has not included.
The model is therefore short by
and that shortfall splits into two pieces with different names. The best energy obtainable from any single product of identical orbitals — the Hartree–Fock limit, also quoted — is a.u. So
The first is what a better shape for the orbital would recover, and it is small: the exponential guess was already close. The second cannot be recovered by any choice of orbital shape whatsoever, because it comes from the electrons avoiding each other instant by instant, which no product of one-electron functions can express. It is called the correlation energy, and recovering it is the subject of most of the companion course Quantum Chemistry. Notice its size: about eV, which is a chemical bond's worth of energy out of eV of binding — small in relative terms and decisive in absolute ones.
Helium has now produced three different “effective nuclear charges”, and none of them is wrong:
, the exponent that minimizes the total energy of the two-electron atom (Example 5.6.1);
, the charge a hydrogenic orbital would need in order to be bound by helium's measured ionization energy;
from Slater's empirical screening rules, fitted to reproduce trends across the periodic table.
Each answers a different question, and the second differs from the first chiefly because removing one electron lets the other relax — an energy difference carries relaxation that an orbital exponent does not. Quoting one where another is meant is the most common way to be wrong by several eV while appearing to use the right idea. Whenever a appears in a table, the question it answers has to be read off with it.
5.11Summary
Fermion states are antisymmetric under exchange (Postulate 5.2.1); Pauli exclusion (Thm. 5.2.1) and the Slater determinant (Def. 5.2.1) follow.
Helium's Hamiltonian, Eq. (5.3), is not separable because of the single term .
Three treatments: independent electrons give eV; first-order perturbation theory, using (Thm. 5.5.1), gives eV; the variational method with a screened exponent gives and eV, against the exact eV.
The variational principle (Thm. 5.6.1) guarantees an upper bound, so a result below the exact energy indicates an error.
Antisymmetry ties spin to spatial symmetry, giving (Thm. 5.7.1). The triplet lies lower by ; for helium's , eV, four orders of magnitude larger than any magnetic spin interaction. This is exchange, and it has no classical analogue.
In heavier atoms each electron moves in a screened central field running from to . Penetration, controlled by the centrifugal barrier, breaks hydrogen's -degeneracy in the order .
This is described by the quantum defect, . For sodium: , , .
Term symbols are ; closed shells contribute and a single hole behaves like a single electron (Thm. 5.9.1), so ionizing a closed shell always gives a doublet.
A doubly excited state can be degenerate with a continuum and decay into it without radiating, a process called autoionization. Helium's at eV has meV, hence fs.
Two electrons in two orbitals have antisymmetric states, sorting as one triplet and three singlets (Example 5.10.1). The same six states reappear as a two-orbital active space in quantum chemistry, as the Hubbard dimer at half filling, and as a two-qubit register.
Optimizing the orbital exponent gains exactly a.u. over first-order perturbation theory, for every (Eq. (5.21) and Fig. 5.5) — of the binding for helium and by .
The model's ionization energy is eV against eV exact. Of the eV shortfall, eV is imperfect orbital shape and eV is correlation, which no product of one-electron functions can supply (Example 5.10.3). That second number is what the companion course Quantum Chemistry spends most of its length recovering.
Helium has at least three defensible effective charges — , and — because each answers a different question (Misreading 5.10.1).
5.12Exercises
The determinant. Write out the Slater determinant for three electrons in orbitals . Verify that interchanging and changes the sign, and that setting gives zero.
The Coulomb integral. Reproduce the proof of Theorem 5.5.1, filling in the two standard integrals used. Then evaluate in eV for , and , and explain physically why the repulsion grows with even though the electrons are being pulled closer together.
The variational calculation.
Derive Eq. (5.10), stating where each of the three terms comes from.
Minimize it to obtain and .
Apply the result to H (, two electrons). What and what energy do you obtain? Compare with the energy of a hydrogen atom plus a free electron ( a.u.) and comment on whether this trial function predicts H to be bound.
Exchange.
Carry out the expansion in the proof of Theorem 5.7.1, showing all four terms explicitly.
Explain in three or four sentences why the spatially antisymmetric state has lower energy, referring to what happens at .
The measured splitting of helium's levels is eV, smaller than the eV of the levels. Using Eq. (5.13), explain why a orbital gives a smaller exchange integral with than a orbital does.
Quantum defects from data. Using eV for sodium and Eq. (5.18), compute and for the state (bound by eV) and the state (bound by eV). Compare with the and values of Example 5.8.1 and state whether behaves as Def. 5.8.1 claims.
Term symbols. Give the ground-state term symbol for He (), Li (), Be () and F (). For fluorine, use Theorem 5.9.1 rather than coupling five electrons.
5.13Project: Screening and the Quantum Defect
The problem. Take the numerical radial solver built in Chapter 4 and insert a screened potential into it. Show that screening breaks the -degeneracy, obtain quantum defects from your own eigenvalues, and compare them with sodium.
A convenient model potential, crude but with the correct limits, is
in atomic units, with for sodium and a parameter setting the core size.
On paper. Verify that Eq. (5.22) satisfies both limits of Eq. (5.15). Sketch it against and , and say where the crossover lies in terms of .
On paper. Before computing, predict the sign of the effect: will move up or down relative to hydrogen's , and will move more or less? Justify the prediction with the centrifugal barrier in two sentences, and record it.
On the computer. Solve Eq. (3.44) with Eq. (5.22) for , starting from . Extract the eigenvalues near and convert each to a quantum defect using Eq. (5.18) with , so that the binding energy is simply .
On the computer. Compare with sodium's measured , , . Tune to fit , then report what the other two do. Does fitting one spoil the others? State which features of the real atom this one-parameter model captures and which it cannot.
On the computer. Plot for in your fitted potential, with the core region shaded. The figure should make the mechanism visible at a glance: the state has weight inside the shaded region and the state almost none. Quantify this by computing for each .
What has to move. One animation: increase the screening smoothly from none (, pure hydrogen) to , showing the three levels starting degenerate and fanning apart with the level falling fastest. That fanning is the breaking of the accidental degeneracy.
The check. At the three levels must be degenerate at a.u. and all three defects must vanish. If they do not, the error is in the solver rather than in the physics, and should be corrected before the screening is switched on.
Be ready to answer. The model has one adjustable parameter and three defects to reproduce. Suppose all three can be fitted by hand. What would that tell you, and what would it not tell you, about whether Eq. (5.22) is a good description of the sodium core?